[an error occurred while processing this directive]
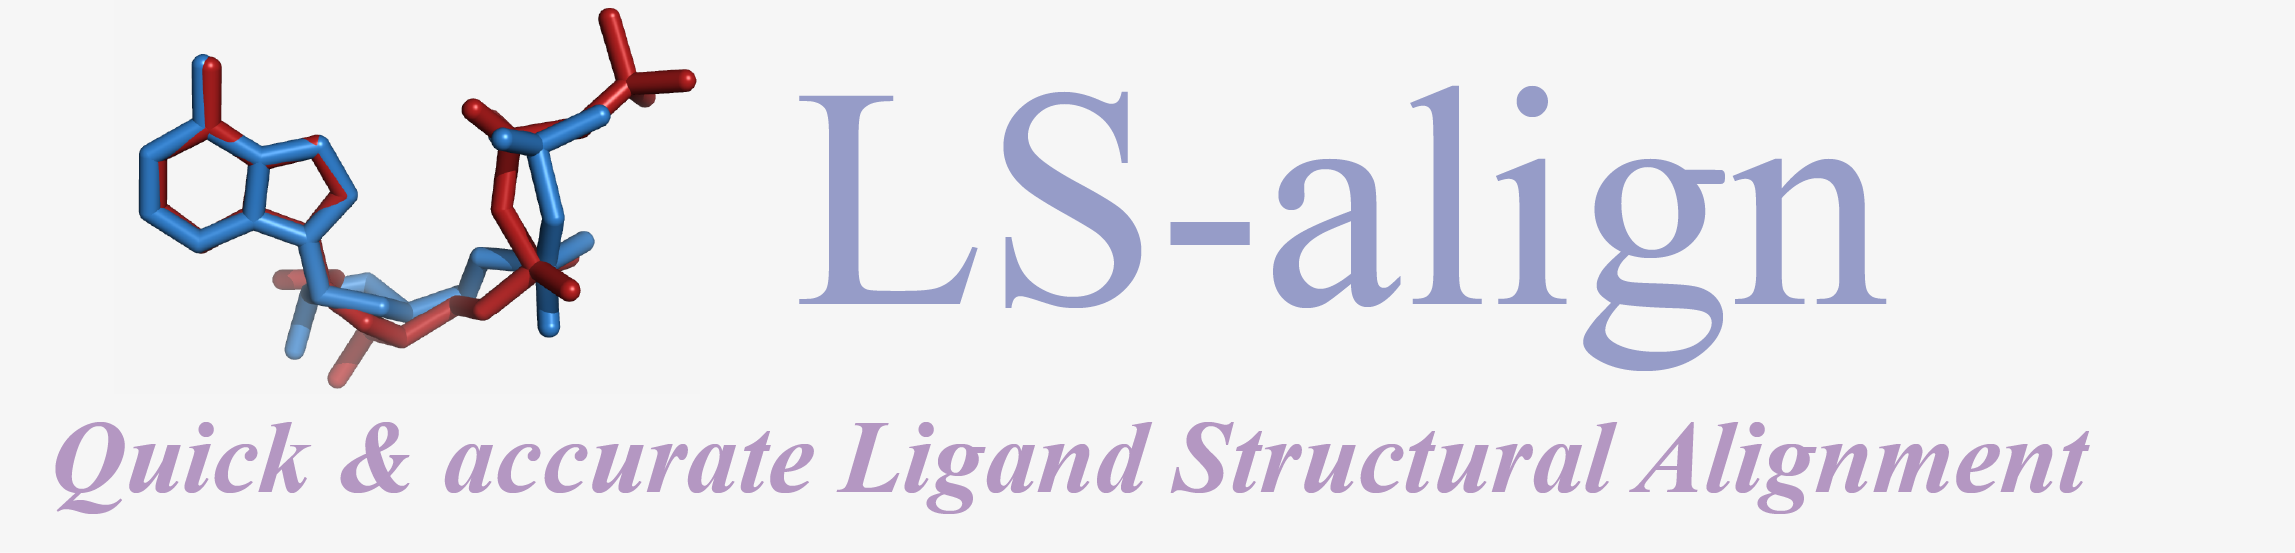
LS-align is an algorithm designed for atom-level structural comparison of ligand molecules.
The target function of LS-align is a combination of inter-atom distance, atom mass, and chemical
bond connections; while the final atom-to-atom alignment is generated by maximizing such
target function through an enhanced-greedy based, iterative heuristic search algorithm.
LS-align program contains two modules:
Rigid-LS-align for rigid-body ligand structure
comparison; and
Flexi-LS-align for flixible structure comparison.
In particular, the
Flexi-LS-align module seeks for optimal alignments of various alternative
conformers of the ligand molecules by rotating flexible bond-angles,
which allows the consideration of binding-induced conformational changes
in the ligand structural comparison and alignment.
[
View an example of output]
[
Download dataset used in LS-align]
LS-align download
-
Click LSalign_src.tar.gz to download the newest version of the source code of LS-align coded in C++. You can compile the program in your Linux computer by:
>tar zxvf LSalign_src.tar.gz
>cd src
>g++ LSalign.cpp -o LSalign
or
>g++ -static -O3 -ffast-math -lm -o LSalign LSalign.cpp
-
Click LSalign.tar.gz to download the executable program for Linux and Windows systems. Nevetheless, it is strongly recommended to download the LS-align source code and compile it on your machine, which gives you higher speed to run the program.
LS-align other information
-
The newest updates of LS-align program:
References:
-
Jun Hu, Zi Liu, Dong-Jun Yu, and Yang Zhang.
LS-align: an atom-level, flexible ligand structural alignment algorithm for high-throughput virtual screening.
Bioinformatics, 34: 2209-2218 (2018).
[Download download the PDF file]
[Support Information]
 umich.edu
| (734) 647-1549 | 100 Washtenaw Avenue, Ann Arbor, MI 48109-2218